VASA-SEQFull-length, plate-based total RNA single-cell sequencing for low cell numbers and FACS-sorted populations
VASA-seq (VASt transcriptome Analysis of single cells by dA-tailing) is a highly sensitive plate-based method that enables you to study full-length RNA and a variety of RNA species; including coding and non-coding.
Exclusively offered by Single Cell Discoveries.

Trusted by

Is VASA-seq the right method for your experiment?
Consider another method when…
- Your samples are non-model species
Probes are currently available for human, mouse, sheep, NHP and rabbit, with other species available on demand. Working with another species? Message us to discuss your project. - You need thousands of cells per sample, or expect many cell types.
Other assays can target anything between 3,000 to 1 000 000 cells per sample at a lower cost per cell. For a standard 10,000-cell human experiment, VASA-Seq may not be suitable. - You cannot FACS-sort your cells.
All our plate-based methods depend on being able to sort. - You do not need single-cell resolution.
Bulk RNA sequencing is compatible with low input material down to hundreds of cells. - You are interested in miRNA.
Although VASA-seq captures non-coding RNAs, miRNAs are generally too short to be reliably detected.
VASA-seq is the right choice when…
- You have a low number of cells.
VASA-seq avoids the 500,000+ cell suspension minimum required by other platforms. - You can FACS-sort your cells.
If your cells cannot be sorted into 384-well plates, VASA-seq is not an option - talk to us about other methods instead. - You need full-length transcripts, non-coding or immature RNA.
When your project demands splicing information, immature RNA, or non-coding RNA, VASA-seq will generate the required output. - You need T-cell receptor repertoire.
At higher VASA-seq sequencing depths, it becomes possible to capture TCR sequences and characterize the TCR repertoire. For higher throughput experiments, 10x Genomics Single Cell Immune Profiling would be better for 5′ gene expression with V(D)J and antigen specificity. - You need hundreds of cells per condition, not thousands.
If your biological question is answered by a few hundred well-characterised cells, this is the method for you. Perfect for prioritizing depth rather than breadth. - Your population of interest is rare, and FACS can gate it.
Sorting puts your target cells of interest in the plate. You spend your sequencing budget only on the population you care about. - Your cells are too large for microfluidic capture.
Whole cardiomyocytes and other large cell types that can clog or bypass a microfluidic chip can be sorted into a well without difficulty. - You want to run a pilot before committing.
Sequence a single plate, look at the data, then decide. See Start with one plate. Plates can be stored at −80 °C and processed in batches later.
Specifications
VASA-seq at a glance
| Format | 384-well cell-capture plate, one cell per well |
|---|---|
| Cells per plate | 376 single cells, plus 8 empty wells as a no-template control |
| Minimum per plate | Half a plate filled with single cells |
| Plates per sample | Two recommended — one processed, one technical backup |
| Cell isolation | FACS, sorted by you or by us |
| Input material | Single Cell Suspension |
| Sample types | Fresh, Cryo-preserved and PFA-fixed |
| Species | Model species |
| Sequencing depth | 150,000 as standard or 1 million for TCR sequencing |
| Sequencing platform | Illumina NovaSeq X Plus |
| Turnaround | 6-10 Weeks |
| Deliverables | FASTQ files, Gene count tables, interactive exploratory data analysis report |
VASA-seq workflow
How a VASA-seq project runs
VASA-seq splits the work between your lab and ours. You do the sorting, because you know your gating strategy and your cells – and we will take it from there.
library prep

Step 1
Consultation
To start, one of our specialist team discusses your biological question, expected cell numbers and experimental set up with you. We act as collaborative partners to support your research.

Step 2
Plates arrive at your lab
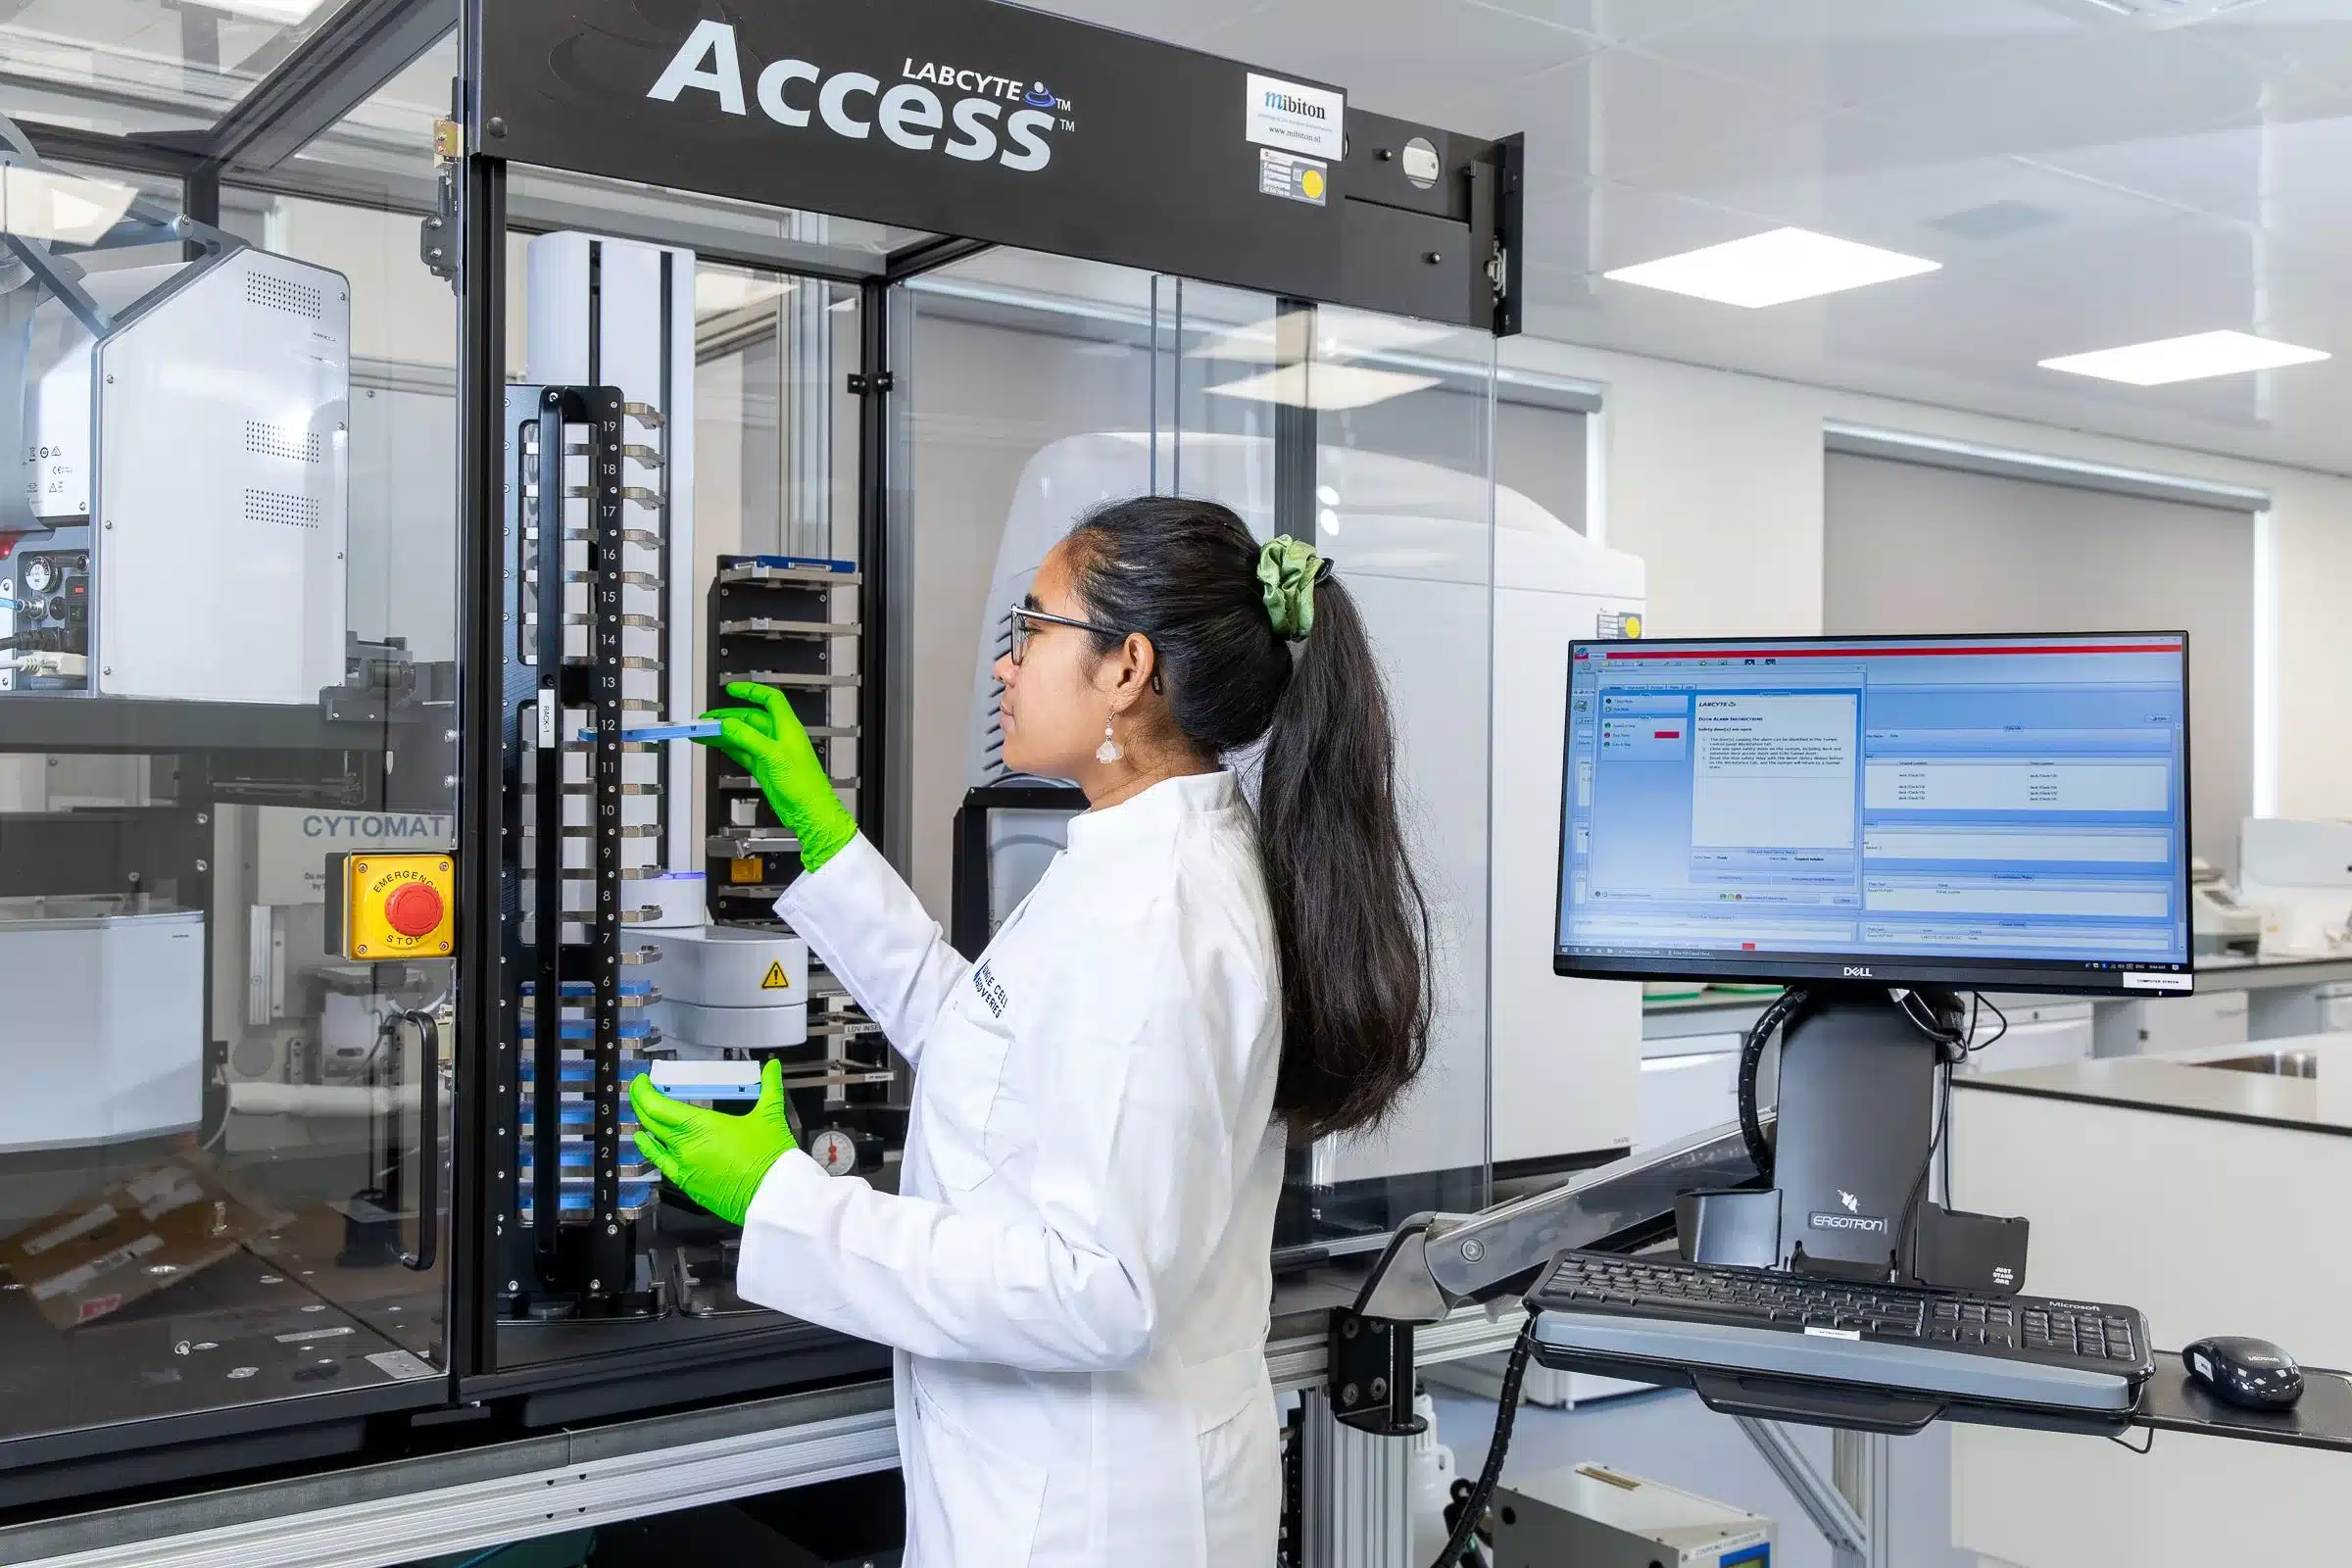
We ship cell-capture plates to you, along with an empty 384-well plate for calibrating your sorter. Every well already contains the barcode and mineral oil. Store the plates are stored at −20 °C until you use them and they can be stored for approximately a year.
Step 3
You dissociate, gate and sort
We will send you detailed instructions on how you sort into the VASA-Seq plate and store the cells before sending to us. Read more on our sample requirements page.
If you do not have access to a sorter, we can offer FACS sorting as a service.

Step 4
You seal and ship the plates back
Seal the plates with the supplied aluminium seals and ship on dry ice with your submission form.

Step 5
Library preparation, sequencing and analysis
Once your samples arrive with us, we begin library prep. RNA from the plate is amplified and prepared as a single sequencing library. We run QC checks along the way, keeping you updated as we go, and finally we sequence on our Illumina NovaSeq X Plus sequencer.

Step 6
Mapping and data analysis
Our data team prepares an exploratory data analysis report and your raw data to send to you.
Deliverables
What you get back
Every VASA-seq project includes:
- Raw sequencing data — FASTQ files
- MultiQC report — raw data statistics generated with FastQC
- Gene count matrices — raw and filtered
velocyto results - Loom file with counts divided in spliced/unspliced/ambiguous per plate - Raw and filtered Seurat objects, so you can pick the analysis up in your own pipeline without reprocessing
- An interactive exploratory analysis report - includes QC metrics and basic filtering, clustering, and differential gene expression analysis, with detailed descriptions of each analysis step and a comprehensive methods section.
- A walkthrough from our data team explaining the quality metrics in your dataset (upon request)
All raw data, results, and reports are processed on EU servers and encrypted at rest using AES-256 (SSE-S3) in Amazon S3 before delivery and archiving.
If you need more than the exploratory report, for example cell type annotation, gene set enrichment analysis, pseudotime analysis, or publication-ready figures; you can book a non-commitment meeting with our data consulting team and discuss your data analysis project
Detecting the total transcriptome
Analyse coding and non-coding RNA
Standard 3′ single-cell RNA-seq can reveal which genes are expressed, but it often provides too little information to understand how transcripts are regulated, processed, or matured. When the biology depends on alternative splicing, isoform usage, non-coding RNAs, RNA velocity, or transcript maturation, a gene-level readout may miss important differences between cells.
VASA-seq captures transcripts across their full length, providing richer information and enabling analyses of splicing and isoform variation, nascent versus mature RNA, and cell-state dynamics. This makes it particularly interesting for studying disease mechanisms, developmental trajectories, and complex cell states where changes in transcript architecture and not just gene expression are biologically important.
VASA-seq allows for many applications involving non-coding and non-polyadenylated RNA types. For example, you can determine the cell cycle stage across developmental phases or between cell populations on a single-cell level based on histone content. The RNA biotypes you can detect include lncRNA, snRNA, snoRNA, and histone genes.
Wondering what technology to use for your project?
Watch this webinar to learn more about choosing the right technology.
Full-length, total RNA single-cell sequencing
How does VASA-seq work?
VASA-seq (Vast Transcriptome Analysis of Single Cells by dA-tailing) is a full-length, total RNA sequencing method that captures both polyadenylated and non-polyadenylated RNA at single-cell resolution. Unlike conventional single-cell RNA sequencing (scRNA-seq) methods that primarily capture poly(A)-tailed mRNA, VASA-seq uses artificial polyadenylation to enable more comprehensive transcriptome profiling.
Following cell lysis and RNA fragmentation, RNA molecules receive an artificial poly(A) tail, allowing oligo-dT primers to initiate reverse transcription. This approach offers several advantages:
- Capture of coding and non-coding RNA: VASA-seq detects a broad range of RNA biotypes, including mRNA, long non-coding RNA (lncRNA), small nucleolar RNA (snoRNA), histone mRNA, and pre-mRNA. This enables researchers to investigate gene expression and RNA regulation beyond conventional mRNA profiling.
- Full-length transcript coverage: Unlike 3′ single-cell RNA-seq methods, VASA-seq provides more uniform coverage across gene bodies. This supports the analysis of transcript structure, alternative splicing, untranslated regions (UTRs), and RNA processing.
- High-resolution transcriptome analysis: By capturing both mature and nascent RNA, VASA-seq provides insights into transcriptional activity, RNA dynamics, and cellular heterogeneity that may be missed by poly(A)-based single-cell sequencing methods.
- Compatibility with Illumina sequencing: VASA-seq libraries can be sequenced using standard Illumina short-read platforms, allowing integration with established bioinformatics workflows for gene expression analysis and transcriptome characterization.
By combining total RNA capture with full-length transcript coverage, VASA-seq offers a more comprehensive view of the single-cell transcriptome, particularly for researchers interested in non-coding RNA, RNA processing, and gene regulation.
Provenance
Where VASA-seq comes from
Developed through a collaboration mainly between researchers at the Hubrecht Institute and Oncode Institute in Utrecht, the Netherlands and Francis Crick Institute, University of Cambridge, in the UK. VASA-seq was introduced in a landmark 2022 publication in Nature Biotechnology, led by Fredrik Salmen and Joachim De Jonghe under the joint supervision of Florian Hollfelder and Alexander van Oudenaarden.
The technology was designed to move beyond traditional poly(A)-based single-cell RNA-seq methods. The original study demonstrated VASA-seq's capabilities by profiling over 30,000 cells from developing mouse embryos, revealing previously undetected cell-type markers, alternative splicing patterns, and RNA dynamics during embryonic development.
Today, VASA-seq still provides researchers with a powerful approach to studying the total transcriptome beyond conventional single-cell RNA sequencing.
2022
First published
240+
Citations
376
Cells per plate
1
Provider worldwide
Pilot
Validate Your VASA-seq Experiment Before Scaling Up
Before scaling up your VASA-seq experiment, start with a pilot to assess sample quality, sequencing performance, and whether your cell type of interest can be detected.
With the raw data, we provide a report to help you evaluate the results. This allows you to assess the number of genes detected per cell, whether your target cell populations can be identified, and whether the expected biological signals are captured.
A pilot experiment helps answer critical questions: Can my cell type of interest be detected? Is the RNA quality sufficient? Can I identify the transcripts and biological differences relevant to my research?
The pilot allows you to validate your experimental design before scaling up to a larger VASA-seq study.
Wondering if this service can
help answer your biological question?
Request a quote today and find out how our expertise can support your research.
What our clients say
"The service of Single Cell Discoveries guarantees a smooth process between customer and company. In case of problems or questions, the scientists reacted competently and immediately, so that answers could be found quickly and flexibly."
"Fantastic job made by a really professional and friendly team. I strongly recommend Single Cell Discoveries for sequencing services."
Recent publications using VASA-seq
Nature
Role of stem-like cells in chemotherapy resistance and relapse in pediatric T-cell acute lymphoblastic leukemia
Case studies
Immune Profiling of atherosclerosis reveals new therapeutic potential
A team of scientists uses 10x Genomics Single Cell Immune Profiling to characterize T cells in the atherosclerotic plaque. Their results redefine atherosclerosis as an autoimmune disease.
Mechanism of disease: a new hypothesis for atherosclerosis
To elucidate atherosclerosis’ mechanism of disease, Sheffield University researchers analyzed the transcriptomes of single endothelial cells.

Drug Discovery
for Precancerous Liver Disease
Here’s the story of how SORT-seq helped advance a novel drug candidate for liver cancer into clinical trials.
Recent blogs related to VASA-seq
Gene set enrichment analysis: From Genes to Biological Processes
Gene set enrichment analysis (GSEA) helps translate gene-level expression changes into broader biological processes. It starts with a ranked list of all tested genes, usually based on a signed statistic…

How to Prevent Single-Cell Sequencing Delays in Pharmaceutical Research
Single-cell sequencing has become an indispensable tool for pharmaceutical R&D. From identifying disease-driving cell populations to discovering predictive biomarkers and evaluating drug response, it provides a level of biological resolution…
Ultra-Low Input RNA-seq: Strategies for Limited or Degraded Samples
RNA sequencing (RNA-seq) has become one of the most powerful tools for studying gene expression. However, not every project starts with abundant, high-quality RNA. Researchers often work with tiny biopsies,…
VASA-seq
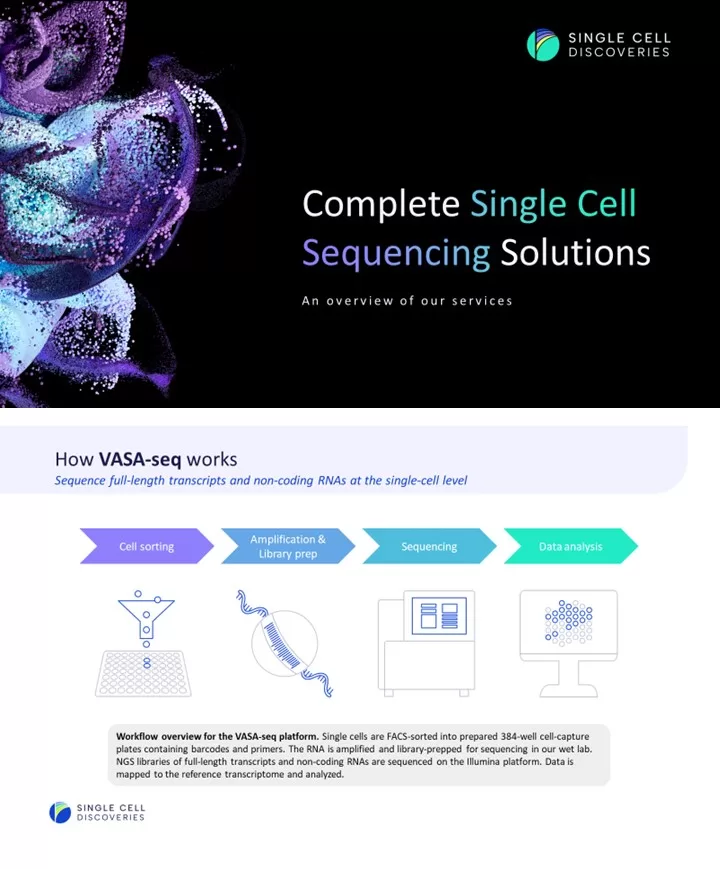
Information Guide
Get an overview of Single Cell Discoveries, VASA-seq, how to get started, and more in our VASA-seq information guide.
FAQ
Probes are currently available for human, mouse, sheep, NHP and rabbit, with other species available on demand. Please contact us if you're interested.
Request a quote
Tell us how many cells you can sort, and we’ll tell you whether VASA-seq fits
Send us your cell numbers. If VASA-seq is the right method, we’ll scope it. If another assay may be more appropriate, we’ll discuss that instead.
Fill out the form for a quote request or book a call with a specialist to find out if VASA-seq is the right solution for your biological question.